Upcoming workshop · 3-Day ADMET & ML Drug Discovery Training
ADMET Profiling, Machine Learning and Docking-Guided Drug Discovery
From molecule to decision: one real EGFR project, step by step
About the workshop
Most drug candidates that fail in development do not fail because they miss their target. They fail because the body never lets enough of the molecule reach that target safely. This three-day workshop teaches ADMET (absorption, distribution, metabolism, excretion and toxicity) as a practical, logical drug-discovery workflow rather than a list of numbers produced by a website.
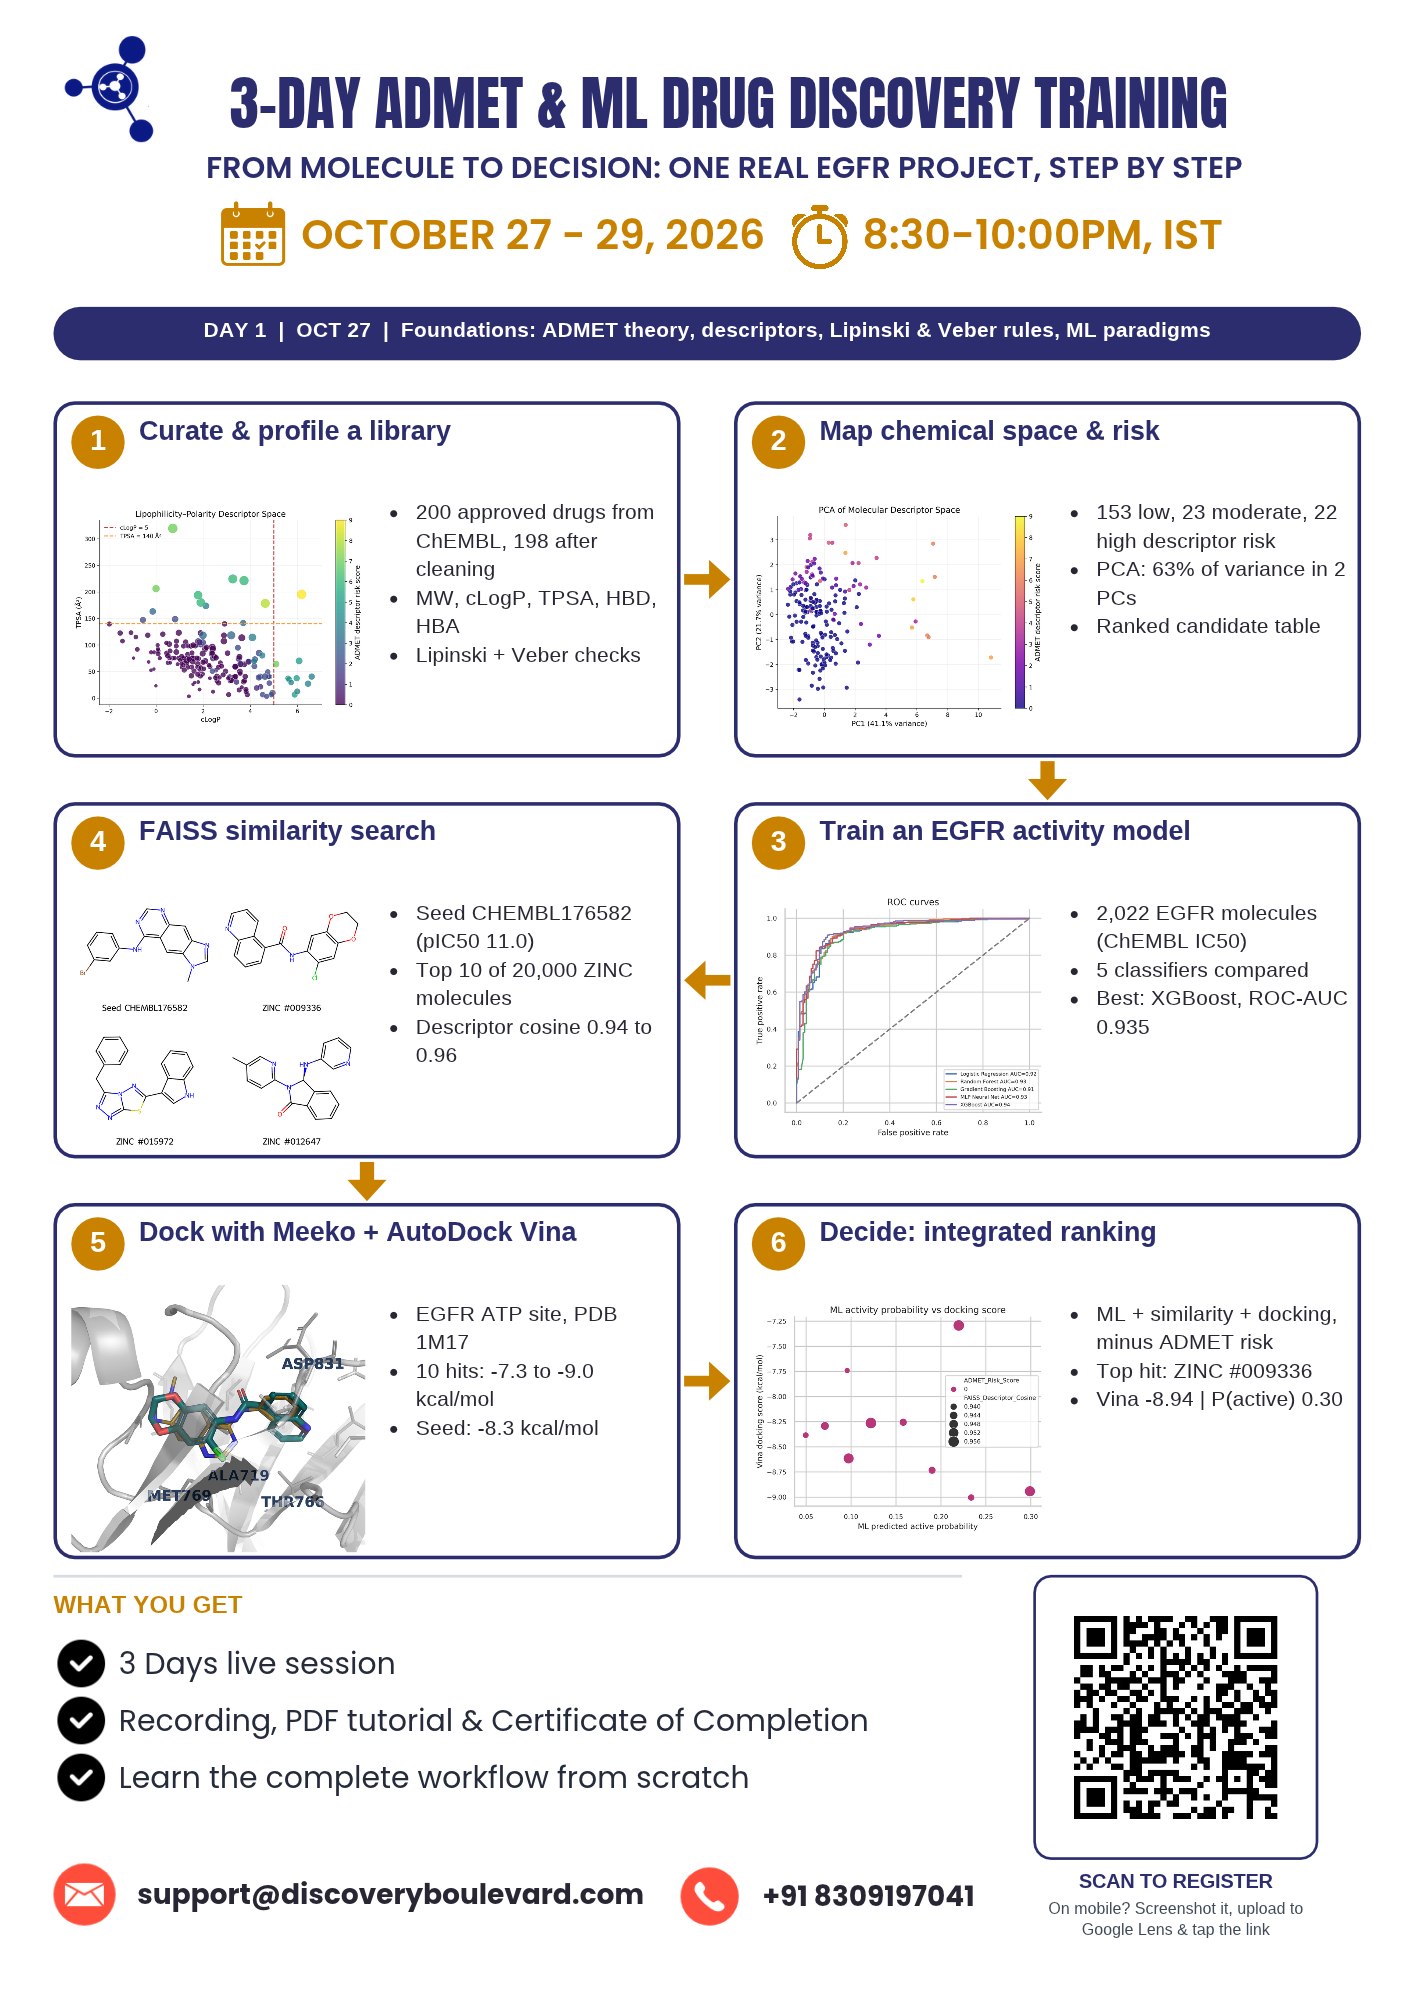
Day 1 builds the biological and computational foundation. Day 2 turns that foundation into an automated ADMET profiling pipeline for a compound library. Day 3 closes the loop with a small end-to-end discovery project: a target-specific machine-learning model, FAISS descriptor-space similarity search, ADMET filtering and docking-supported validation against EGFR kinase (PDB 1M17).
Every participant runs the full workflow in their own browser through Google Colab. No prior programming experience is required.
- Live online
- Google Colab notebooks
- Session recordings
- PDF tutorial
- Certificate of Completion
Curriculum
What the workshop covers
Three evenings that take one real EGFR project from molecular structure to a developability decision.
- Day 1 · Tue 27 Oct
ADMET Theory and Machine Learning-Based Prediction
The biological and computational foundation of ADMET, before any automated analysis.
Why candidates fail
- Why potent molecules fail: Fialuridine and Torcetrapib
- ADMET as the life story of a drug
- Molecular properties versus developability
Key parameters and rules
- Solubility, Caco-2, PAMPA, P-gp efflux, plasma protein binding and BBB
- CYP metabolism, clearance, half-life, hERG, Ames and DILI
- MW, cLogP, TPSA, HBD, HBA and rotatable bonds
- Lipinski and Veber rules as warning lights
Machine learning paradigms
- Supervised endpoint prediction
- Unsupervised chemical-space exploration
- Reinforcement learning for optimization
- Day 2 · Wed 28 Oct
Automated ADMET Profiling
Turning a molecular library into an interpretable ADMET profile ranked by developability risk.
Library and curation
- 200 reference molecules from ChEMBL, or your own SMILES file
- SMILES validation and canonicalization with RDKit
- Salt stripping and duplicate removal
Visualization and profiling
- 2D structure grids and MMFF-optimized 3D conformers
- Interactive py3Dmol views
- Core descriptors and automated Lipinski and Veber evaluation
Decisions
- cLogP versus TPSA maps and descriptor heatmaps
- PCA of chemical space
- ADMET risk flags and a risk-ranked candidate table
- Day 3 · Thu 29 Oct
ML-Guided Discovery and Molecular Docking
Connecting ADMET profiling with target-specific prediction, similarity search and docking.
Activity models
- EGFR IC50 data from ChEMBL converted to pIC50
- Classifiers including random forest and XGBoost
- ROC-AUC, F1, precision, recall and confusion matrices
Similarity search
- FAISS search for the 10 molecules closest to a known active
- ML activity probability and ADMET risk for each hit
Docking and ranking
- Ligand preparation with Meeko
- Docking into EGFR (PDB 1M17) with AutoDock Vina
- Pose comparison and an integrated final ranking
What you receive
- Three ready-to-run Google Colab notebooks
- PDF tutorial
- Recorded videos of all sessions
- Certificate of Completion
- Practical outputs: descriptor tables, plots, rankings, docking scores and poses
- A full structure-to-decision workflow you can reuse on your own targets
Who it is for
- M.Sc., M.Pharm and Ph.D. students
- Researchers in bioinformatics, computational chemistry and drug discovery
- Medicinal chemists who want a fast, interpretable computational triage
- Faculty looking for a ready ADMET and ML practical, and beginners: no programming experience needed
Register
Fees & registration
₹1,200 / $20
Rupees for participants in India, US dollars for international participants.
1. Fill in the registration form
Go to the form2. Pay the workshop fee
After paying, email the transaction screenshot to support@discoveryboulevard.com.
Good to know
- • All three days are paid sessions.
- • The certificate, PDF tutorial and recorded videos are provided to registered paid participants.
- • Bring a laptop with an updated Chrome or Edge browser and a Google account for Google Colab.
- • Basic chemistry and biology knowledge is recommended.
- • The registration fee is non-refundable and non-transferable.
Registration and queries
Registration form
Form not loading? Open it in a new tab.